The EU Clinical Trials Regulation (Regulation (EU) No 536/2014) changed the way clinical trial applications are submitted in the EU and EEA. Since February 2023, all clinical trial applications must have been submitted via the Clinical Trials Information System (CTIS) portal.
The transition deadline ended in January 2025. By that date, clinical trials approved under the previous Directive had to be transferred to CTIS if still ongoing.
Below we have summarised the most important CTIS-related points from the perspective of investigator-initiated studies.
WHAT IS THE CTIS PORTAL?
CTIS is a system developed by the European Medicines Agency (EMA) for managing and overseeing clinical trials. The implementation of CTIS is a key part of the EU Clinical Trials Regulation, which aims to harmonise the regulatory requirements and enable a single submission route for multi-country clinical trials.
- The CTIS portal consists of two separate workspaces:
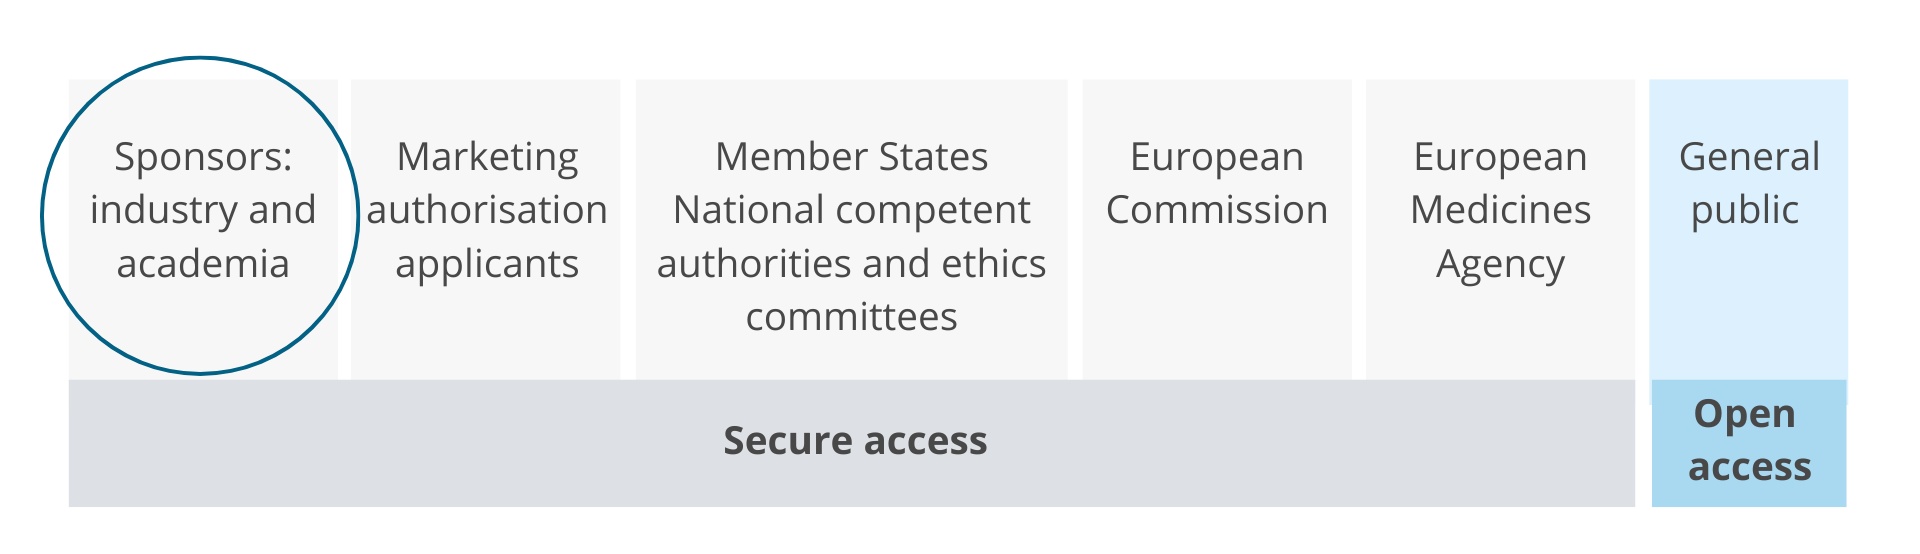
- The sponsor workspace, intended for sponsors. Here, the sponsor can prepare, compile and submit clinical trial data to Member States for assessment. Tasks can also be delegated to Contract Research Organisations (CROs), such as Medfiles.
- The authority workspace, intended for national competent authorities, ethics committees, the European Commission and the EMA. This is where Member States and the Commission assess clinical trial applications.
- In addition, there is a public website that is accessible to patients, healthcare professionals, researchers, CROs, the media and the general public.
WHY THE CTIS PORTAL WAS ESTABLISHED?
The CTIS portal was designed to simplify and streamline the clinical trial application and approval process. Its goal is to offer a centralised and harmonised platform for submitting, assessing and managing clinical trial applications across all EU and EEA countries. The system facilitates communication and collaboration between national competent authorities, ethics committees and sponsors.
- Applications can be submitted simultaneously to both authorities and ethics committees across the EU and EEA.
- Both submission and trial-related reporting are managed centrally via a single portal.
- CTIS increases the transparency of clinical trials – for all parties involved, including participants.
It is important to note that the portal must also be used for clinical trial applications for investigator-initiated studies, even if the trial takes place in a single country and at a single site – unless it is considered a non-interventional study.
Free Guide: Quick tips for successful CTIS submissions
The CTIS portal brings new responsibilities and requirements and raises questions. We created a practical guide on how to successfully submit your clinical trial applications. The information is especially targeted at academic researchers conducting clinical trials in Finland.
HOW TO GET STARTED WITH THE CTIS PORTAL?
Before you can start, you need to check the following:
1. Do you have an EMA account?
- To use the CTIS sponsor workspace, you must have an active EMA user account. If you already use other EMA systems (e.g. OMS, IRIS, EudraVigilance), your existing credentials can be used.
- If you do not yet have an active EMA account, you must register via self-registration.
2. Is your organisation registered in OMS?
- Organisations must be registered in EMA’s Organisation Management Service (OMS).
- Each organisation (e.g. university hospital, university, or CRO) has one Organisation ID, but may have multiple Location IDs. You should check this with your institution’s research or administrative department.
3. Have you registered a sponsor administrator?
- The sponsor administrator must be registered via EMA account management. Depending on the submission strategy your organisation chooses, you may select either a trial-centric model or an organisation-centric model.
- Instructions for trial-centric submission are available on the authority’s website (PDF, in English).
- The trial-centric model is intended especially for small organisations such as academic research groups conducting trials occasionally. Please check with your organisation to confirm the appropriate approach. The system will also indicate if a CT Administrator has already been assigned.
- The CT Administrator role is automatically assigned to the user who initiates a new clinical trial application in CTIS, but it is strongly recommended that at least one back-up CT Administrator is assigned.
- Users may also be granted the roles of Viewer, Preparer or Submitter. These are managed by the CT Administrator.
WHEN TO START using CTIS?
- The transition period under the EU Clinical Trials Regulation (536/2014) ended on 30 January 2025. Clinical trials authorised under the previous Directive are no longer considered legally valid if they were not submitted as transition applications to CTIS before this date.
- If a legacy study is still ongoing in 2025 and was not transitioned, a full new application must be submitted to CTIS as soon as possible to regularise the situation.
- This requirement also applies to investigator-initiated studies and to studies in the follow-up phase, where subjects are still being contacted (e.g. by phone).
- If you are planning a new trial, start preparing for CTIS submission already during the development of your study protocol.
- Please also ensure that your protocol complies with the Regulation. It is recommended to use the template available on the authority’s website under “Protocol and Consent Form”, where regulatory requirements are pre-included.
- Ensure in advance that you have all documents required for the application, including documentation related to authorised investigational medicinal products (IMPs), if relevant to your study.
CTIS enhances collaboration and communication between stakeholders involved in the clinical trial process, contributing to more consistent and timely application reviews. The portal also improves transparency by granting public access to key information on authorised clinical trials, including summaries of protocols and trial results.
CTIS also introduces new responsibilities and obligations for investigators. If you are unsure how to begin or need support with a specific part of the process, Medfiles’ Clinical Trials Team is happy to support you. We provide practical, tailored guidance throughout your clinical trial – from start to submission and beyond.
You may also be interested in: